Mastering MedTech Regulations 2026: A Comprehensive Guide for US Healthcare Professionals on FDA Updates
The landscape of medical technology (MedTech) is in a constant state of flux, driven by rapid innovation, evolving scientific understanding, and an unwavering commitment to patient safety. For healthcare professionals in the United States, staying abreast of the latest regulatory changes is not merely a recommendation but a critical imperative. As we approach 2026, the U.S. Food and Drug Administration (FDA) continues to refine and introduce new guidelines that will significantly impact the development, approval, and post-market surveillance of medical devices. This comprehensive guide aims to demystify the upcoming MedTech Regulations 2026, providing a clear roadmap for healthcare professionals to navigate these changes effectively.
Anúncios
Anúncios
Understanding these regulations is paramount for several reasons. Firstly, compliance ensures the legal and ethical operation of healthcare facilities and the appropriate use of medical devices. Non-compliance can lead to severe penalties, including fines, product recalls, and reputational damage. Secondly, a deep understanding of these regulations fosters innovation. By knowing the boundaries and requirements, developers can design devices that not only meet clinical needs but also satisfy regulatory scrutiny from the outset. Finally, and perhaps most importantly, these regulations are designed to protect patients. They ensure that medical devices are safe, effective, and perform as intended, thereby improving patient outcomes and public health.
This article will delve into the key areas of focus for the FDA in 2026, including significant updates to pre-market approval processes, enhanced post-market surveillance requirements, the growing emphasis on cybersecurity in medical devices, and the intricate world of artificial intelligence (AI) and machine learning (ML) in MedTech. We will also explore the implications for various stakeholders, from device manufacturers and healthcare providers to researchers and patients. Our goal is to equip you with the knowledge and insights needed to confidently adapt to the evolving regulatory environment and continue to deliver high-quality, safe, and innovative healthcare.
The Evolving Landscape of MedTech Regulations 2026
The FDA’s approach to regulating medical devices is dynamic, reflecting advancements in technology and lessons learned from past experiences. The MedTech Regulations 2026 are not a sudden overhaul but rather a continuation and refinement of ongoing efforts to modernize the regulatory framework. Key drivers for these changes include the increasing complexity of medical devices, the integration of software and AI, the growing threat of cyberattacks, and the global interconnectedness of the medical device supply chain.
Modernization of Pre-Market Review Pathways
One of the primary areas of focus for the FDA in 2026 is the continuous modernization of its pre-market review pathways. This includes the 510(k) pre-market notification, Pre-Market Approval (PMA), and De Novo classification requests. The FDA is actively seeking to streamline these processes without compromising safety and effectiveness. This often involves leveraging real-world evidence (RWE) and real-world data (RWD) more extensively to support regulatory submissions. Healthcare professionals involved in clinical trials and data collection will find their roles increasingly critical in generating the robust evidence required for regulatory success.
The FDA’s push for a ‘least burdensome’ approach aims to reduce unnecessary regulatory hurdles while maintaining high standards. However, ‘least burdensome’ does not mean ‘less rigorous.’ Instead, it emphasizes efficiency and the use of appropriate scientific methods to gather sufficient evidence. For manufacturers, this translates to a need for early engagement with the FDA and a clear understanding of the specific data requirements for their device type. Healthcare providers, particularly those involved in early device adoption or clinical research, should be aware of these evolving evidence requirements as they can influence which devices becomes available and how quickly.
Enhanced Post-Market Surveillance and Real-World Evidence
Beyond pre-market approval, the FDA is placing a significant emphasis on enhanced post-market surveillance. The goal is to detect safety signals and performance issues more quickly once a device is in widespread use. This involves a multi-faceted approach, including:
- National Evaluation System for health Technology (NEST): NEST continues to evolve as a centralized platform for gathering and analyzing real-world data from various sources, including electronic health records, claims data, and patient registries. Healthcare organizations are encouraged to contribute to and utilize NEST to enhance the collective understanding of device performance in diverse patient populations.
- Unique Device Identification (UDI) System: The UDI system remains a cornerstone of post-market surveillance. By 2026, compliance with UDI labeling and database submission requirements is expected to be fully entrenched across almost all device classes. Healthcare professionals involved in inventory management, purchasing, and clinical documentation must ensure their systems are capable of capturing and utilizing UDI information to facilitate traceability and recall management.
- Patient and Clinician Reported Outcomes (PROs/CROs): The FDA is increasingly recognizing the value of direct patient and clinician feedback in assessing device performance and safety. Expect more emphasis on structured methods for collecting and integrating PROs and CROs into post-market surveillance programs.
For healthcare professionals, this means an increased responsibility in reporting adverse events accurately and promptly, participating in registries, and potentially contributing to studies that generate real-world evidence. The data you generate in your daily practice can directly influence regulatory decisions and improve device safety for future patients.
The Rise of Digital Health and AI/ML in MedTech Regulations 2026
Perhaps no area of MedTech is evolving as rapidly as digital health, encompassing mobile medical applications, wearable sensors, and artificial intelligence/machine learning (AI/ML) driven diagnostics and therapeutics. The MedTech Regulations 2026 will significantly address how these innovative technologies are regulated.
Software as a Medical Device (SaMD)
The FDA has been developing a regulatory framework for Software as a Medical Device (SaMD) for several years, and 2026 will see further maturation of these guidelines. SaMD refers to software intended to be used for medical purposes without being part of a hardware medical device. Examples include software that analyzes medical images for diagnostic purposes or algorithms that predict disease risk based on patient data.
Key aspects of SaMD regulation include:
- Risk-Based Classification: SaMD is classified based on its intended use and the impact of its information on patient care, ranging from low-risk wellness apps to high-risk diagnostic or therapeutic software.
- Pre-Certification (Pre-Cert) Program: While still under pilot, the Pre-Cert program aims to evaluate the quality and organizational excellence of SaMD developers rather than solely focusing on individual products. If successful, this could streamline the review process for companies with a proven track record of developing safe and effective software.
- Validation and Performance: Demonstrating the clinical validity and analytical validity of SaMD is crucial. This involves rigorous testing to ensure the software accurately processes data and produces clinically meaningful results.
Healthcare professionals using SaMD must understand its regulatory status, how it was validated, and any limitations on its use. Education and training on these digital tools will become increasingly important to ensure their appropriate and safe integration into clinical practice.
Artificial Intelligence and Machine Learning (AI/ML) in Medical Devices
AI/ML represents a paradigm shift in medical device functionality, offering capabilities like adaptive learning and continuous improvement. However, these very characteristics pose unique regulatory challenges. The FDA is actively developing a regulatory framework for AI/ML-driven medical devices, focusing on:
- Transparency and Explainability: Understanding how AI algorithms arrive at their conclusions (interpretability) and the factors influencing their performance is critical.
- Bias Mitigation: Ensuring AI/ML algorithms are trained on diverse datasets to prevent biases that could lead to health disparities.
- Algorithm Change Protocol: For ‘locked’ algorithms, traditional regulatory pathways suffice. However, for ‘adaptive’ or ‘continuously learning’ algorithms, the FDA is exploring mechanisms to ensure ongoing safety and effectiveness post-market, potentially through pre-defined change control plans.
- Data Management and Governance: The quality, integrity, and security of the data used to train and operate AI/ML models are paramount.
Healthcare professionals using AI/ML-powered devices need to be aware of their limitations, understand how to interpret their outputs, and recognize situations where human oversight is indispensable. Continuous education on AI ethics and application in clinical settings will be vital.
Cybersecurity: A Non-Negotiable Aspect of MedTech Regulations 2026
In an increasingly interconnected healthcare ecosystem, medical devices are prime targets for cyberattacks. A compromised device can not only expose sensitive patient data but also directly endanger patient safety by altering device functionality. Recognizing this critical threat, cybersecurity has become a cornerstone of MedTech Regulations 2026.
Pre-Market Cybersecurity Requirements
The FDA now mandates robust cybersecurity documentation as part of pre-market submissions for many medical devices. This includes:
- Cybersecurity Risk Management: Manufacturers must demonstrate a comprehensive approach to identifying, assessing, and mitigating cybersecurity risks throughout the device’s lifecycle.
- Software Bill of Materials (SBOM): Providing an SBOM, which lists all software components used in a device, helps identify vulnerabilities and facilitates rapid responses to newly discovered threats.
- Security Controls: Devices must incorporate appropriate security controls, such as authentication, authorization, data encryption, and secure updates.
- Testing and Validation: Manufacturers are expected to conduct rigorous cybersecurity testing, including penetration testing and vulnerability assessments.
Post-Market Cybersecurity Management
- Vulnerability Management: Manufacturers must have processes in place to monitor for new vulnerabilities, assess their impact, and issue patches or updates in a timely manner.
- Coordinated Vulnerability Disclosure (CVD): Encouraging collaboration between manufacturers, researchers, and government agencies to disclose and address vulnerabilities responsibly.
- Incident Response Plans: Having clear plans for responding to cybersecurity incidents, including communication with affected users and regulatory bodies.
For healthcare professionals and organizations, this translates to a heightened need for vigilance. IT and clinical engineering departments must work closely to implement robust network security, regularly update device software, and train staff on cybersecurity best practices. Understanding the cybersecurity posture of medical devices before procurement is also becoming a key decision factor.
Global Harmonization and Supply Chain Resilience
The global nature of medical device manufacturing and distribution means that US regulations are often influenced by and, in turn, influence international standards. The FDA actively participates in harmonization efforts with bodies like the International Medical Device Regulators Forum (IMDRF) to streamline regulatory processes and ensure consistent safety standards worldwide. This is particularly relevant for devices manufactured abroad and imported into the US, or for US-made devices intended for export.
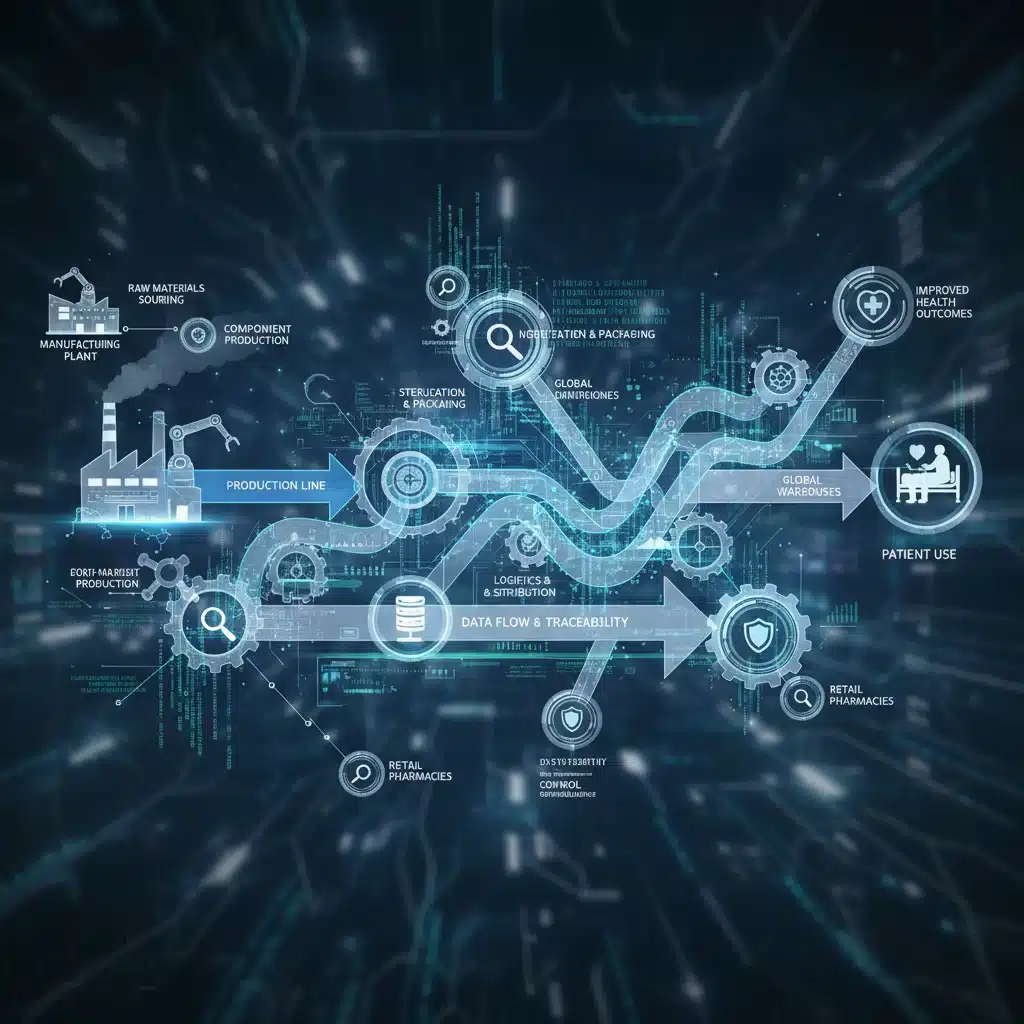
Furthermore, recent global events have highlighted the fragility of supply chains. The MedTech Regulations 2026 will likely continue to emphasize supply chain resilience, requiring manufacturers to demonstrate robust strategies for mitigating disruptions, ensuring the quality of components from suppliers, and maintaining transparency throughout the product’s journey from raw material to patient.
Healthcare professionals need to be aware of the origins of their medical devices and the supply chain practices of their vendors. This knowledge contributes to risk management and ensures access to essential medical technologies even during periods of global instability.
Implications for US Healthcare Professionals
The evolving MedTech Regulations 2026 have profound implications for virtually every healthcare professional in the US. Here’s a breakdown of how these changes might affect different roles:
- Physicians and Surgeons: You will need to stay informed about the latest approved devices, especially those incorporating AI/ML, and understand their validated uses and limitations. Your role in reporting adverse events and contributing to real-world evidence will be increasingly critical.
- Nurses and Allied Health Professionals: As frontline users of medical devices, your adherence to proper device operation, maintenance, and reporting protocols is essential for patient safety and regulatory compliance. Training on new digital health tools and cybersecurity hygiene will be paramount.
- Hospital Administrators and IT/Clinical Engineering Staff: You bear the primary responsibility for ensuring that devices procured and used within your facility meet regulatory standards, are securely integrated into your network, and that staff are adequately trained. Developing robust cybersecurity policies and incident response plans is non-negotiable.
- Researchers and Clinical Trial Coordinators: Your work in generating high-quality clinical evidence and real-world data will directly support regulatory submissions and post-market surveillance. Adherence to ethical guidelines and data integrity standards is more important than ever.
- Purchasing and Procurement Teams: You will need to conduct thorough due diligence on vendors, assessing not only product efficacy and cost but also regulatory compliance, cybersecurity posture, and supply chain resilience.
Continuous education and professional development will be key to adapting to these changes. Healthcare organizations should invest in robust training programs to ensure their staff are well-versed in the latest regulatory requirements and best practices for using modern medical technologies.
Preparing for MedTech Regulations 2026: A Strategic Approach
Proactive preparation is essential to navigate the MedTech Regulations 2026 successfully. Here are some strategic steps healthcare professionals and organizations can take:
- Stay Informed Through Official Channels: Regularly monitor FDA guidance documents, public workshops, and advisory committee meetings. Subscribe to FDA newsletters and reputable industry publications.
- Invest in Continuous Education and Training: Develop comprehensive training programs for all staff involved in the use, procurement, and management of medical devices. Focus on new technologies like AI/ML, cybersecurity protocols, and updated reporting requirements.
- Strengthen Cybersecurity Infrastructure: Conduct regular cybersecurity audits, implement robust network segmentation, ensure timely software updates for medical devices, and train staff on identifying and reporting potential threats.
- Enhance Data Governance and Management: Ensure that systems for collecting, storing, and analyzing patient data and device performance data are robust, secure, and compliant with privacy regulations (e.g., HIPAA) and FDA requirements. Leverage UDI for improved traceability.
- Fostering Collaboration: Encourage cross-functional collaboration between clinical, IT, legal, and administrative departments within your organization. Engage with industry associations, manufacturers, and regulatory experts to share best practices and insights.
- Develop a Proactive Compliance Strategy: Don’t wait for enforcement actions. Regularly review your internal policies and procedures to ensure they align with the latest FDA guidelines. Consider mock audits or internal assessments to identify and address gaps.
- Engage with Manufacturers: Maintain open communication with medical device manufacturers regarding their regulatory compliance efforts, cybersecurity updates, and post-market surveillance activities.
Conclusion: Embracing the Future of MedTech with Confidence
The year 2026 marks another significant chapter in the evolution of MedTech regulations. While the changes may seem daunting, they represent the FDA’s ongoing commitment to fostering innovation while safeguarding public health. For US healthcare professionals, embracing these new guidelines is not just about compliance; it’s about leading the charge in delivering safer, more effective, and technologically advanced patient care.
By proactively understanding and integrating the nuances of the MedTech Regulations 2026, healthcare organizations can transform potential challenges into opportunities. Opportunities to enhance patient safety, optimize operational efficiency, and drive forward the adoption of groundbreaking medical technologies responsibly. The future of healthcare is inextricably linked with the responsible evolution of medical technology, and your role in navigating this complex, yet exciting, landscape is more crucial than ever.
Stay informed, stay proactive, and continue to prioritize patient well-being as we collectively shape the future of medical innovation under the vigilant eye of the FDA.